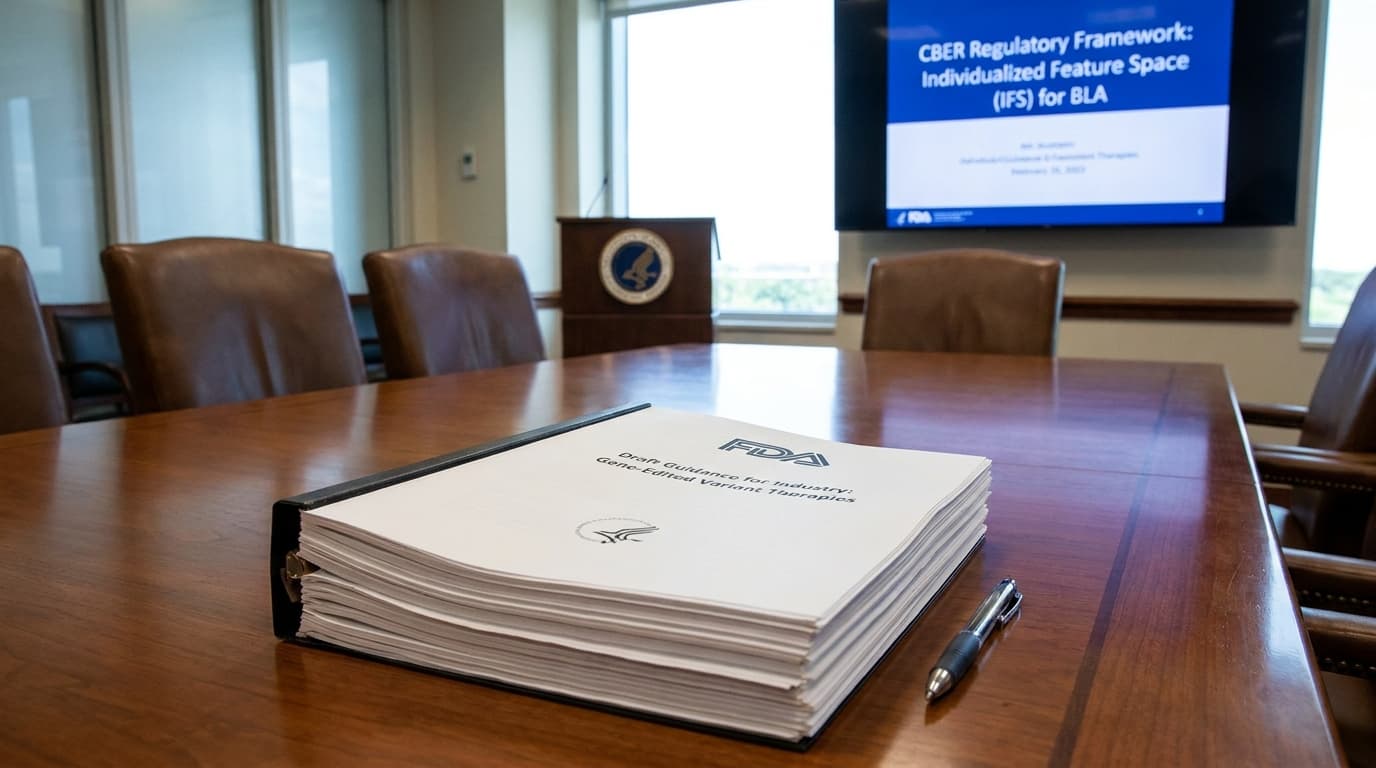
The FDA has released draft guidance that lays out how developers of gene-edited therapies could seek approval for multiple mutation-specific products through a single biologics license application. The move gives concrete form to a regulatory idea former biologics chief Peter Marks described in 2024: treating closely related CRISPR-based medicines as platform variants rather than wholly separate products.
For the field, the signal is less about a single document than about market structure. If the approach holds, companies building editing platforms around common delivery systems and manufacturing processes may gain an advantage over one-off programs, because each added variant could face a lighter path than a stand-alone application.
The policy framework
Current Center for Biologics Evaluation and Research Director Karim Mikhail told BioSpace the guidance offers a path for approvals to happen in a more expeditious way. The draft guidance describes how one regulatory application could cover gene-editing therapies addressing various disease-causing mutations, with the goal of enabling faster access for patients in need.
The central concept is the individualized feature space, or IFS. According to the draft, the IFS defines the parameters for variations on a product that can be grouped together as part of a set of customized products.
Andrew Byrnes, director of the Division of Gene Therapy 1 at CBER, framed the rationale during an FDA webinar by comparing the human genome to an instruction manual with three billion letters, where a genetic disease can result from a single typo and different patients with the same disease may have different typos.
BMO Capital Markets said the core premise is to create faster and more efficient access with a consistent IFS that serves as an anchor for a single BLA across all variations, where each additional variant would need to produce predictable safety and activity.
The road here
The framework has been in discussion for years. At the American Society of Gene & Cell Therapy’s 27th annual conference in May 2024, Peter Marks said one advantage of CRISPR-based therapies is that companies could make many different products targeting different mutations in the same gene and that the FDA could theoretically create regulatory processes to simplify approvals for those groups.
Marks argued then that many CRISPR constructs aimed at the liver for different conditions could be “99.9% identical,” making a platform-style manufacturing and regulatory model plausible. Less than a year after those remarks, he was ousted as director of CBER. Two and a half years later, the agency has now advanced the concept into draft guidance.
H.C. Wainwright characterized the move as the FDA turning its plausible mechanism pathway into a “BLA blueprint.”
Who stands to benefit
BioSpace identified Beam Therapeutics and Prime Medicine as at least two biotechs with development programs that could benefit from the approach.
Beam uses base editing, which changes single DNA letters directly without cutting both strands of the double helix. The company is developing precision genetic medicines for alpha-1 antitrypsin deficiency, glycogen storage disease Ia and phenylketonuria, or PKU. For PKU, Beam’s BEAM-304 is in a Phase 1/2 trial and is designed to correct mutations in the phenylalanine hydroxylase gene. H.C. Wainwright said the potential permutations of BEAM-304’s suite of guide molecules fit the FDA’s draft framework, and called Beam’s inclusion of multiple PKU-causing mutations in its Phase 1/2 study a landmark development for the field.
Prime Medicine, which develops medicines based on prime editing, could also benefit. BMO highlighted PM577a, a one-time treatment in a Phase 1/2 trial for Wilson disease variants caused by an H1069Q mutation in the ATP7B gene. BMO said follow-on editors could address other ATP7B variants using largely common delivery and editing components, and that the draft guidance could give Prime an important regulatory tailwind if subsequent mutation-specific editors can rely primarily on laboratory evidence once a shared platform and clinically predictive acceptance criteria are established.
The broader implication is that as researchers keep finding new mutations, regulatory efficiency may become a competitive asset in gene editing alongside biology and delivery.